|
Research Description
Our major research topics focus on
bioinformatics and biological problem study. What I have been doing is to
develop useful bioinformatics tools and novel methodologies not only for
service to a wide range of biologists but also for ourselves to study
important biological problems as well as to answer interesting biological
questions. Although I am a computer scientist by training, I firmly believe
that studying biological problems and questions through collaboration with
biologists can definitely enhance our understanding of what problems and
questions are important and interesting to the biology community and what
kinds of new bioinformatics tools are urgently needed for biologists.
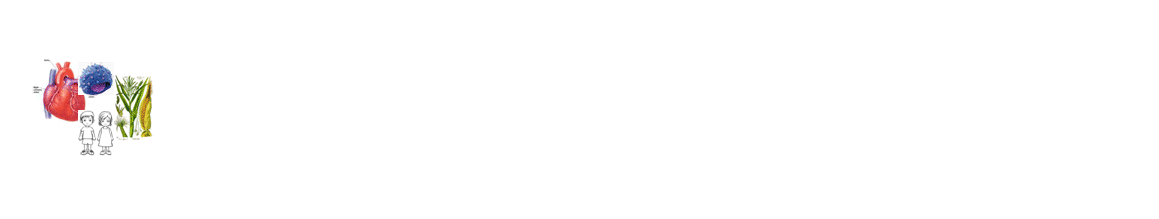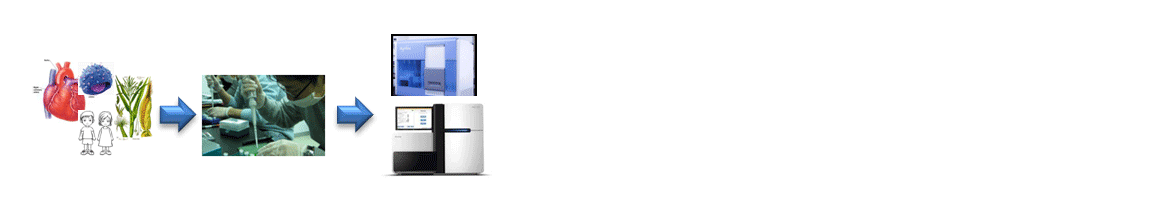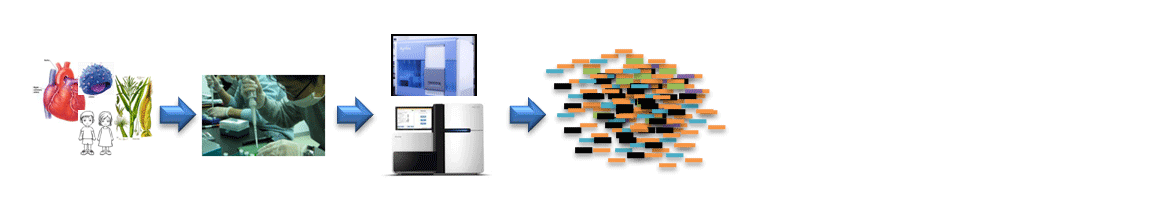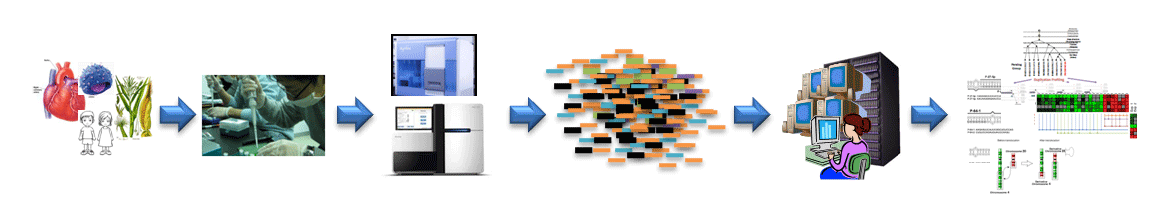
Currently, our lab focuses on using the
next-generation sequencing (NGS) technology to study the genomics and transcriptomics related topics including: (a) gene
regulation and molecular evolution of Kranz anatomy
formation during C4 plant development, (b) microRNAs in diseases and B cell
differentiation, and (c) development of short read sequence assembler and
related analysis tools. In what follows, I will introduce these topics and
the collaborators in detail.
(a) Gene regulation and molecular
evolution of Kranz anatomy formation
during C4 plant development
Most plants can
be divided into two types: C3 and C4. C4 plants, such as maize, have more
efficient photosynthesis than C3 plants and can survive better in more
extreme environments, such as high temperatures and arid lands. However,
several food plants, such as rice or wheat, are all C3 plants; so their growth environments and areas are very
limited. If we can increase the photosynthesis efficiency or even construct
the C4 photosynthesis pathway to these C3 food plants, their productivity
can be definitely increased and planting environments can be extended to
more areas. It will be greatly helpful to solve the food crisis for Third
World countries. However, the regulatory network of photosynthesis is very
complex. Particularly, C4 plants require the coordination of
the mesophyll and bundle sheath cells, called Kranz leaf anatomy, to confer high rates of
photosynthesis. Moreover, how many genes are involved in the Kranzanatomy formation and how they regulate the
formation during the early leaf development are
largely unknown. In 2010, Academician Dr. Wen-Hsiung
Li formed a group to study this topic. The participants are from different
institutions in Academia Sinica and National Chiayi University, Taiwan and their professional
specialties include plant photosynthesis, plant physiology, gene
transformation, molecular evolution, bioinformatics, and computational
biology. We want to study gene regulatory differences between C3 and C4
photosynthesis pathways and the Kranz anatomy
formation during the early leaf development. In this project, our lab is
responsible for processing and analyzing transcriptomic deep
sequencing data, predicting key regulators of Kranz anatomy
formation and C4 photosynthesis pathway, and reconstructing gene regulatory
networks of C3 and C4 photosynthesis pathways.
(b) MicroRNAs co-targeting in diseases
and B cell differentiation
MicroRNAs (miRNAs) play an important role in development, cell
differentiation, and diseases. Most related studies only focused on
studying regulatory mechanisms of individual miRNAs
and single target genes. But most gene regulations are so complex and may
compensate each other in vivo. That is, individual target genes
could be regulated by a set of miRNAs and several
miRNAs may co-target a common target gene.
However, only few studies on miRNA co-targeting
have been reported in literature. In this study, we are working on
development of a systematic approach to integrate the experimental data,
acquired from NGS and other platforms, and the information, from databases
and literature, to predict miRNA co-targeting
networks in diseases and cell differentiation. The topics we want to study
include breast cancer metastasis (with Dr. Yu, Alice Lin-Tsing, GRC, Academia Sinica, Taiwan), B cell differentiation (with Dr. Lin, Kuo-I,
GRC, Academia Sinica, Taiwan), and cardiac hypertrophy (with Dr.
Chen, Chien-Chang, IBMS,
Academia Sinica, Taiwan). We have sequenced small RNA samples
collected from these collaborators' laboratories. We start to analyze the
sequenced raw read data and will predict potential co-targeting pathways
and finally validate by experiment. We expect that our results can increase
the knowledge of miRNA regulations in different
diseases and cell differentiation.
|
(c) Development of short read sequence assembler
and related analysis tools
Short read sequencing (SRS)
platforms, including Illumina Genome Analyzer II and HiSeq 2000 and AppliedBiosystems SOLiD with high throughput and relatively low
cost, have been widely used for genomic andtranscriptomic studies
in recent years. However, in contrast to the Sanger technique, which
produces reads of ~1000 bp, short read
sequencers produce shorter but a much greater number of reads, thus
posing new challenges to many computational issues, such as de
novo genome and transcriptome assemblies.
A set of state-of-the-art approaches for processing SRS data were
developed while the read lengths of SRS were from 40 bp to 76 bp.
However, the read length and throughput have been greatly improved. The
read length byIllumina Genome
Analyzer IIx (GAIIx),
for example, has been increased to ~150 bp,
and the throughput has been increased from 12 Gb to 95 Gb per
run. Although longer read length and higher numbers of reads provide more
information for data processing, the computer memory requirements and
execution times usually also increase with read length and total read
count. In this topic, we have focused on developing a new de novo assembly
algorithm to assemble a genome using Illumina short
reads alone. The prototype has been almost completed and achieves better
memory usage, speed, and contig quality than many current
methods. Next, we will focus on developing effective and efficient
algorithms to identify genome structural variations and the breakpoints
by Illumina short read data directly.
|
|
|
|
|
|
|
|